|
|
Diamond 5Following the recent release of Diamond version 5, a new Diamond version 5.1 will become available, offering new functions and enhanced commands for working with Metal Organic Frameworks (MOFs):�nbsp;Improved selection of molecular or polymeric fragments as well as selection of symmetry-related molecules or fragments. �nbsp;Copy Style and Paste Style will be enhanced, so you can copy the design properties of multiple selected atoms and bonds to another (selected) fragment in one step. �nbsp;Creation and application of design schemes from/to selected atoms, bonds, and polyhedra, so you can quickly define designs for frequently used organic linkers etc. �nbsp;Improved adjustment of structure picture to ensure that atoms or bonds selected in the tables become visible in the structure picture. The update to version 5.1 is free-of-charge for all customers of version 5 (and version 4). The release of version 5.1 is planned for the first half of 2024. |
 |
Match! 4We are currently working on the final steps for the new Match! version 4. The main improvement will be the so-called "profile-fitting search-match" (PFSM), a powerful, innovative alternative to the proven peak-based search-match functionality. The new function fits the profile calculated from each candidate reference entry to the experimental profile on the fly, so that the sometimes ambiguous peak searching is no longer required for qualitative analysis.While the success rate of this new approach is definitely higher than the "classical" (peak-based) approach (especially with minor or trace phases), there is certainly one main drawback: The calculation time is significantly longer. In order to compensate for this to a certain degree, the profile-fitting search-match runs in parallel on all CPU cores. All Match! customers whose update permission time has not expired yet will be able to update to the new version free-of-charge. We currently plan to release Match! 4 in the first half of 2024. |
 |
Endeavour 2We have started working on a new Endeavour version 2. The main new improvement will be the option to run multiple calculations with different seed-values in parallel on multiple CPU cores.This new functionality will allow you to easily distribute the different seed values of a structure solution calculation to the available CPU cores. By doing so you can use the full potential of your CPU and make structure solution calculations much faster (depending on the number of available CPU cores, of course). In addition, the solution of crystal structures from single crystallite electron diffraction data will be improved. Finally, an option will be added to "watch" the update of a certain crystal structure data file. Each time it gets updated e.g. by an external application (like some molecular modeling program or similar), Endeavour will update the crystal structure picture automatically, using the current Autobuild Settings. The release of version 2 is estimated for the second half of 2024. |
New Match! version 3.16 (Build 283)
December 18, 2023
The new Match! version 3.16 Build 283 is a maintenance release in which all currently known bugs have been fixed.
Here is the full list of improvements in the new version:
- An issue has been resolved that might cause an error message like "You are not entitled to use this more recent version of Match!" after (re-)installation, even though the installed version is still within the range of your update permission time.
- [Windows] On Windows systems it was not possible to use the mouse wheel, e.g. to correct the zero point error or for zooming.
- A bug affecting the import/indexing of old PDF-2 databases (prior to 2005) has been fixed.
- [macOS] Due to a bug, Match! could not be installed on macOS versions earlier than 10.15 "Catalina", even though the (correct) minimum macOS version 10.13 "High Sierra" is mentioned in the system requirements.
- Some minor bugs have also been fixed.
- New release of the COD reference database
六个产品
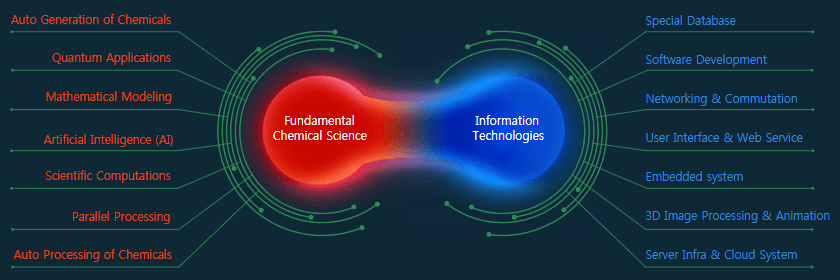
从化学科学的角度来看åQŒ我们的ç‰Ҏ®Š½Ž—法可以快速,自动地生成大量化学物质。量子力学用于基¼‹€分析åQŒ我们先˜q›的数学建模和ä�h工智能(AIåQ‰技术可提供高质量的化学信息。我们特ŒDŠ的计算和处理技术ä‹Éòq¶行˜q�行数千台计½Ž—机成äؓ可能åQŒ这些计½Ž—机可以快速自动处理数十亿¿U�化学药品ã€?/span>
我们的特ŒDŠ信息技术可以提供广泛的在线服务。我们的数据库技术专门用于复杂的¿U‘学信息åQŒ能够搜索数十亿¿U�化学物质的¾l“构。我们的软äšg技术可以处理复杂的¿U‘学现象åQŒ例å¦?/span>3D化学囑փ�处理和动甅R€‚我们还拥有与硬件相关的技术,例如¾|‘络åQŒ嵌入式¾pȝ»ŸåQŒ服务器基础¾l“构和云¾pȝ»Ÿ构徏ã€?/span>
核心技术图
我们的核心技术是融合
基础化学¿U‘学和信息技术的¾l“果ã€?/span>
|
Software News and Updates |
|
|
CRYSPLOT is a publicly accessible web-based tool (http://crysplot.crystalsolutions.eu) to visualize physico-chemical properties of periodic systems (i.e. crystals, surfaces and polymers) as computed with the CRYSTAL code. It has been designed as a very intuitive graphical tool, a low entry-level interface, to all types of users, from beginners to experts, and purposes, from education to research. |
|
|
|
|
|
|
SPECIAL OFFER for CRYSTAL09/06/03 USERS CRYSTAL09/06/03 Academic users that purchase the new release CRYSTAL17 within December 31, 2019 will be entitled to a 30% discount on the basic license. |
|
|
|
|
Remember! CRYSTAL14 academic users are entitled of 50% discount!
|
|
|
|
Crystal Solutions is a joint project between the Theroretical Chemistry Group of University of Turin and Aethia Srl for CRYSTAL distribution. |
------------------------------------------------------------------------------------------------------------------------------
CRYSTAL on the cover
The performance of the code in terms of parallel efficiency, speed-up and memory distribution of themassively parallel version of the CRYSTAL17program has recently been presented on a paper entitled �/span>Large Scale Condensed Matter DFT Simulations: Performance and Capabilities of the CRYSTAL Code�[J. Chem. Theory Comput., 13, 5019-5027 (2017)].
The code is shown to run and scale efficiently up to 32,000 cores on HPC architectures for the study of systems containing up to 14,000 atoms per cell.
----------------------------------------------------------------------------------------------------------------------------------------------------
CRYSPLOT is an online web-oriented tool to visualize computed properties of periodic systems totally free for CRYSTAL users.
It is targeted for plotting properties of crystalline solids computed with CRYSTAL code and in particular, one can represent band structure and density of states, among many others. It permits the modification and customization of plots to meet the standards required for scientific graphics.
--------------------------------------------------------------------------------------------------------------------------
NEW TUTORIALS ONLINE!
Have a look at the restyled website of CRYSTAL online Tutorials with an enhanced readability, more content, and a fancier look!
Tutorials have been also updated to cover the new features of CRYSTAL17.
---------------------------------------------------------------------------------------------------------------------------------------------------
HOW TO GET CRYSTAL17
Ordering, download, installation, advice about hardware and other activities related to the use and development of the code are available through the Crystal Solutionswebsite.
--------------------------------------------------------------------------------------------------------------------------------------------------
Remember! CRYSTAL14 academic users are entitled of 50% discount!
]]>
|
|
Details about the new functions and bug fixes that have been implemented are available at
http://www.crystalimpact.com/news/20170922a.htm
Issues when updating from Match! 3.4.x to 3.5.x
-----------------------------------------------
A few customers have reported issues (like a dialog showing a wrong message that the update subscription time has passed) when updating their Match! version 3.4.x (or ealier) installation to one of the more recent versions 3.5 or 3.5.1. These issues were caused either by runtime library incompatibilities and/or license file directory issues.
If you are updating from a Match! version 3.4.x (or earlier) and you are encountering any difficulties when installing or running the new version, please follow these instructions:
First of all, please copy your license file "yourlicense.lic" from your current/old Match! program directory (e.g. C:\Program Files (x86)\Match3") to a safe place (e.g. a temporary directory). Afterwards, uninstall Match!. Finally, manually remove all Match! program directories that might still exist, using the Windows Explorer.
You are now ready to download and run the corresponding new version installer from
http://crystalimpact.de/match/download.htm#download
Once you have installed the new version, please move your license file "yourlicense.lic" from the temporary directory back into the new Match! program directory. You should now be able to run Match! as usual.
]]>
|
|